An in-depth exploration of the Molecular Hamiltonian, covering its key components, applications in quantum chemistry, and analytical methods in molecular physics.
Introduction to the Molecular Hamiltonian
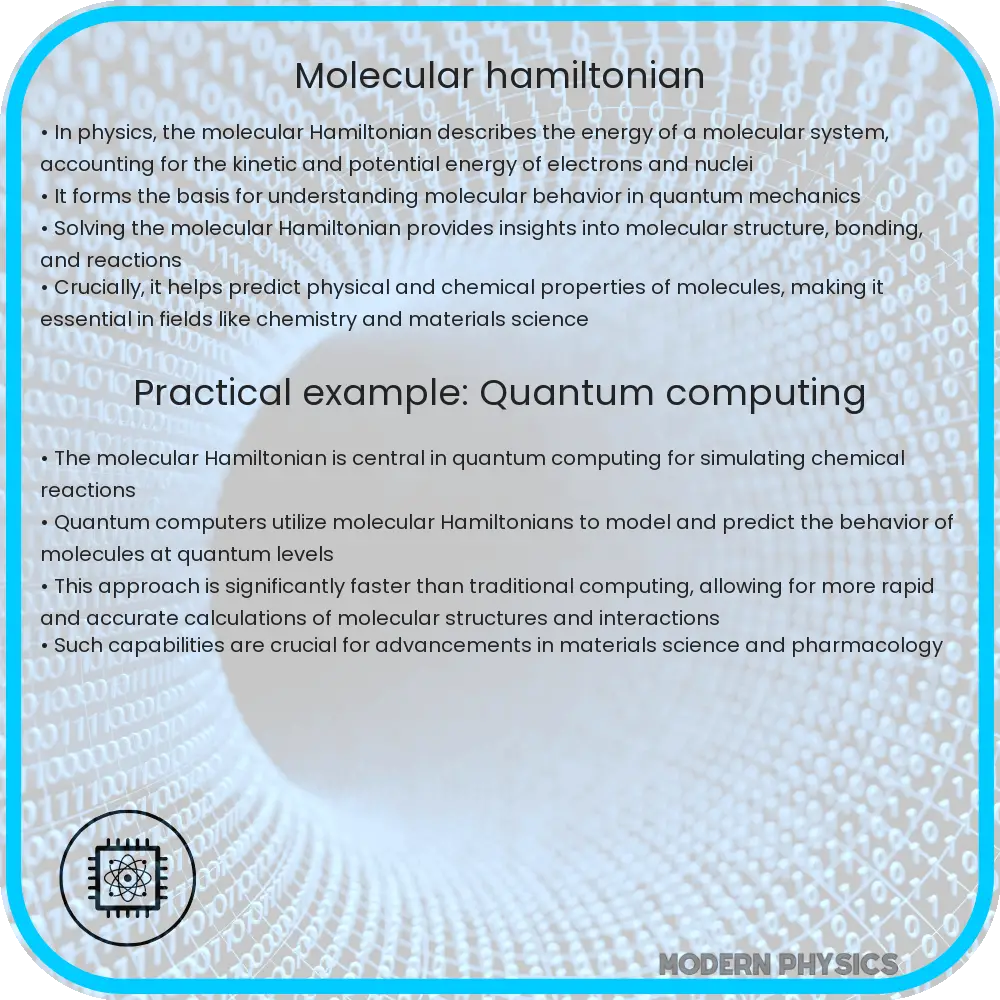
The Molecular Hamiltonian is a fundamental concept in quantum mechanics, particularly in the study of molecular systems. It describes the total energy of a molecule in terms of the kinetic energies of electrons and nuclei, as well as the potential energies resulting from their interactions. Understanding the molecular Hamiltonian is crucial for exploring molecular structure, dynamics, and reactivity in chemical physics.
Key Components of the Molecular Hamiltonian
The molecular Hamiltonian, often denoted as Ĥ, can be expressed by the equation:
\[ \hat{H} = \hat{T}_n + \hat{T}_e + \hat{V}_{nn} + \hat{V}_{ne} + \hat{V}_{ee} \]
where:
- \(\hat{T}_n\) represents the kinetic energy of the nuclei,
- \(\hat{T}_e\) represents the kinetic energy of the electrons,
- \(\hat{V}_{nn}\) is the potential energy due to nuclear-nuclear repulsion,
- \(\hat{V}_{ne}\) is the potential energy due to nuclear-electron attraction, and
- \(\hat{V}_{ee}\) is the potential energy due to electron-electron repulsion.
This breakdown is the starting point for many sophisticated computational methods used to solve the Schrödinger equation for molecules, which is a central problem in theoretical chemistry and molecular physics.
Applications in Quantum Chemistry
The molecular Hamiltonian plays a pivotal role in quantum chemistry, where it is used to predict molecular properties and behaviors. It serves as the basis for:
- Molecular orbital theory, which helps understand the distribution of electrons in a molecule and the chemical bonds formation,
- Electronic structure methods like Hartree-Fock and post-Hartree-Fock, which provide approximations to solve the Schrödinger equation,
- Density Functional Theory (DFT), which uses electron density rather than wavefunction to calculate molecular properties,
- Spectroscopy simulations, which help in predicting the results of experimental techniques that probe the electronic structures of molecules.
Each of these applications has the goal of expanding our understanding of molecular systems, enhancing the accuracy of predictions about chemical properties and the results of chemical reactions, and facilitating the design of new molecules and materials with specific desired properties.
Analysis of the Molecular Hamiltonian
To fully employ the molecular Hamiltonian, scientists often simplify the full Hamiltonian through various approximations. The Born-Oppenheimer approximation is a primary example, where the nuclei’s movement is separated from the electrons due to their mass disparity. This simplification leads to more manageable calculations and has been instrumental in the advancement of computational chemistry.
Further, symmetry considerations can significantly reduce the computational workload. By identifying symmetry operations that leave the Hamiltonian invariant, one can decompose the problem into smaller, more tractable parts. This approach is pivotal in both reducing the computational cost and increasing the feasibility of calculations for larger molecules.
Challenges and Limitations
While the Molecular Hamiltonian provides a comprehensive framework for studying molecules, there are inherent challenges and limitations in its application. One major challenge is the computational power required to solve the Schrödinger equation accurately for large molecules. Even with approximations like the Born-Oppenheimer and using symmetry, the computational effort can be immense, especially when dealing with complex systems.
Furthermore, the accuracy of some methods, such as Hartree-Fock, can be limited due to the approximation of electron correlation. Advanced techniques like Configuration Interaction (CI) and Coupled Cluster (CC) methods are often necessary to achieve greater accuracy, which further increases the computational demand.
Future Prospects
The ongoing developments in computational power and algorithms continuously expand the capabilities of quantum chemistry. The adoption of quantum computing holds particular promise for tackling the Molecular Hamiltonian’s complexities more efficiently. Quantum algorithms have the potential to perform calculations that are currently impractical with classical computers, paving the way for new discoveries in molecular science.
In addition, the integration of machine learning techniques with traditional quantum chemistry methods is opening new avenues for predicting molecular behavior and designing materials with unprecedented precision. These advancements are set to revolutionize the field, making the deep insights provided by the Molecular Hamiltonian even more accessible and actionable.
Conclusion
The Molecular Hamiltonian is more than just a mathematical expression; it is a gateway to understanding the fundamental properties of molecules and their interactions. At its core, it enables scientists and engineers to delve deeper into the quantum world, offering predictions and insights that guide experimental chemistry and material science. Whether it’s through improving existing computational methods or pioneering new ones like quantum computing, the exploration of the Molecular Hamiltonian continues to be at the forefront of scientific advancement, pushing the boundaries of what we know about the molecular world.
As we advance, the application of these principles will undoubtedly yield more sophisticated materials and drugs, improved energy capture and storage technologies, and ultimately, a better understanding of the fabric of our universe. The journey through quantum chemistry, propelled by the Molecular Hamiltonian, promises to be as exciting as it is challenging.